Thursday, April 16, 2009


Lumberjacks and obesity genes
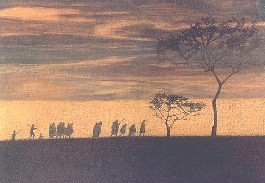
 Thought I'd post on this one, because the title caught my eye, and because I was keen on finding a good lumberjack picture. They look for association between variants in 18 genes and obesity related phenotypes. I'm a bit perplexed by the choice of genes...
- Association study between candidate genes and obesity-related phenotypes using a sample of lumberjacks.
- Chamberland A, Tremblay N, Audet M, Gilbert B, Pérusse L, Vohl MC, Laprise C.
INTRODUCTION: Complex traits such as obesity are modulated by genetic and environmental factors and lead to varied clinical presentations.The aim of this study was to investigate associations between candidate genes and obesity-related phenotypes using a sample of 252 lumberjacks issued from a founder population and sharing a common and circumscribed environment. METHODS: Thirty-seven variants in 18 genes were genotyped. The restriction fragment length polymorphism method and the template-directed dye-terminator incorporation assay with fluorescence polarization detection were employed for the genotyping assays. Multivariate logistic regression models were built in order to calculate the relative odds of exhibiting obesity-related phenotypes associated with the presence of the studied polymorphism. Among them, 21 single nucleotide polymorphisms were tested for associations with obesity phenotypes. RESULTS: Significant associations were found between carriers of the minor alleles of APOE-epsilon2, FABP2-A54T, UCP1-L229M, LPL-HindIII, LPL-S447X and LPL-T1973C, patients bearing a combination of LPL-D9N, LPL-N291S and LPL-P207L and obesity-related phenotypes. CONCLUSION: The present results suggest that a particular population such as lumberjacks, sharing the same environment, could help target genes involved in complex traits.
Sunday, April 12, 2009
Answer to my question (sorta): heritability of melanoma
I wondered about the heritability of tanning response in a recent post about a GWAS for tanning response. Albeit not a direct answer, according to this paper below, there is a relatively high degree of heritability (~55%) for variation in development of melanoma:
Shekar SN, Duffy DL, Youl P, Baxter AJ, Kvaskoff M, Whiteman DC, Green AC, Hughes MC, Hayward NK, Coates M, Martin NG.
J Invest Dermatol. 2009 Apr 9.
Shekar SN, Duffy DL, Youl P, Baxter AJ, Kvaskoff M, Whiteman DC, Green AC, Hughes MC, Hayward NK, Coates M, Martin NG.
J Invest Dermatol. 2009 Apr 9.
Abstract: Melanoma runs within families, but this may be due to either shared genetic or shared environmental influences within those families. The concordance between pairs of non-identical twins compared to that between identical twins can be used to determine whether familial aggregation is due to genetic or environmental factors. Mandatory reporting of melanoma cases in the state of Queensland yielded approximately 12,000 cases between 1982 and 1990. Twins in this study and from the adjacent state of New South Wales (125 pairs in total) were used to partition variation in liability to melanoma into genetic and environmental factors. Identical twins were more concordant for melanoma (4 of 27 pairs) than non-identical twins (3 of 98 pairs; P-value approximately 0.04). Identical co-twins of affected individuals were 9.8 times more likely to be affected than by chance. However, non-identical co-twins of affected individuals were only 1.8 times more likely to be affected than by chance. An MZ:DZ recurrence risk ratio of 5.6 suggests that some of the genetic influences on melanoma are due to epistatic (gene-gene) interactions. Using these data and population prevalences, it was estimated that 55% of the variation in liability to melanoma is due to genetic influences.
Friday, April 10, 2009
Evolutionary medicine gaining ground
 Description of Evolutionary Medicine meeting in the current Science:
Apparently there is a growing trend of teaching medicine from an evolutionary perspective in medical schools. The figure here shows the increase in connections between ecology & evolution and Medicine as reflected in citations (look at connections between the two reddish dots).
Some interesting parts from Elizabeth Pennisi's article:
For instance, anthropologist Kathleen Barnes of Johns Hopkins University has evidence that for some asthmatics, this overly energetic inflammatory response may be a holdover from the body's successes in coping with parasitic disease.Also, in this issue, from the same meeting, a piece about schizophrenia and autism by Constance Holden, arguing for the evolutionary connection between these two conditions (I think that an equally appropriate spectrum is: William's syndrome-Autism)
...With genomic data in hand, "medical students are much more equipped to understand the connections between all organisms,"...
Barnes and her colleagues have found that asthma is associated with the defective Duffy gene in populations in Brazil, Columbia, and the Caribbean whose recent African ancestors lived where malaria was endemic.
At the Sackler Colloquium on Evolution in Health and Medicine held last week at the National Academy of Sciences (NAS) in Washington, D.C., evolutionary geneticist Bernard Crespi of Simon Fraser University in Burnaby, Canada, threw some evolutionary firepower at the question. He proposes that both schizophrenia and autism are disorders of the "social brain"--but at opposite ends of the same spectrum.
A number of studies have shown some overlap in genomic "hot spots" for CNVs in schizophrenia and autism, with, in some cases, deletions in one condition just where there are duplications for the other
That would fit with their theory that psychotic disorders--including not only schizophrenia but also bipolar disorder and some major depression--result from "overdevelopment" of the social brain, and autism spectrum disorders reflect underdevelopment of that brain. Many scientists believe socialization is the main force behind the rapid expansion of human brains, said Crespi, pointing out that in primates the size of the cortex increases with size of social groups. The components of the social brain, according to Crespi, include language, self-awareness, "social emotions" such as pride and guilt, logical thinking, pursuit of goals, and awareness of the mental states of others.
Monday, April 06, 2009
Utility of GWAS prostate cancer risk alleles in other populations
Waters KM, Le Marchand L, Kolonel LN, Monroe KR, Stram DO, Henderson BE, Haiman CA.
Cancer Epidemiol Biomarkers Prev. 2009 Mar 24. [Epub ahead of print]
Abstract: Genome-wide association studies have identified multiple common alleles associated with prostate cancer risk in populations of European ancestry. Testing these variants in other populations is needed to assess the generalizability of the associations and may guide fine-mapping efforts. We examined 13 of these risk variants in a multiethnic sample of 2,768 incident prostate cancer cases and 2,359 controls from the Multiethnic Cohort (African Americans, European Americans, Latinos, Japanese Americans, and Native Hawaiians). We estimated ethnic-specific and pooled odds ratios and tested for ethnic heterogeneity of effects using logistic regression. In ethnic-pooled analyses, 12 of the 13 variants were positively associated with risk, with statistically significant associations (P less than 0.05) noted with six variants: JAZF1, rs10486567 [odds ratio (OR), 1.23; 95% confidence interval (95% CI, 1.12-1.35); Xp11.2, rs5945572 (OR, 1.31; 95% CI, 1.13-1.51); HNF1B, rs4430796 (OR, 1.15; 95% CI, 1.06-1.25); MSMB, rs10993994 (OR, 1.13; 95% CI, 1.04-1.23); 11q13.2, rs7931342 (OR, 1.13; 95% CI, 1.03-1.23); 3p12.1, rs2660753 (OR, 1.11; 95% CI, 1.01-1.21); SLC22A3, rs9364554 (OR, 1.10; 95% CI, 1.00-1.21); CTBP2, rs12769019 (OR, 1.11; 95% CI, 0.99-1.25); HNF1B, rs11649743 (OR, 1.10; 95% CI, 0.99-1.22); EHBP1, rs721048 (OR, 1.08; 95% CI, 0.94-1.25); KLK2/3, rs2735839 (OR, 1.06; 95% CI, 0.97-1.16); 17q24.3, rs1859962 (OR, 1.04; 95% CI, 0.96-1.13); and LMTK2, rs6465657 (OR, 0.99; 95% CI, 0.89-1.09). Significant ethnic heterogeneity of effects was noted for four variants (EHBP1, Phet = 3.9 x 10(-3); 11q13, Phet = 0.023; HNF1B (rs4430796), Phet = 0.026; and KLK2/3, Phet = 2.0 x 10(-3)). Although power was limited in some ethnic/racial groups due to variation in sample size and allele frequencies, these findings suggest that a large fraction of prostate cancer variants identified in populations of European ancestry are global markers of risk. For many of these regions, fine-mapping in non-European samples may help localize causal alleles and better determine their contribution to prostate cancer risk in the population.
Friday, April 03, 2009
GWAS deluge
I wonder how exactly they operationalize this phenotype...I don't have full text access, but I'll assume they control for constitutive skin color. I still think this is a bit weird. Is there a heritable component to skin burning that is really independent of constitutive skin color?
Genome-Wide Association Study of Tanning Phenotype in a Population of European Ancestry.
Nan H, Kraft P, Qureshi AA, Guo Q, Chen C, Hankinson SE, Hu FB, Thomas G, Hoover RN, Chanock S, Hunter DJ, Han J.
J Invest Dermatol. 2009 Apr 2
Genome-Wide Association Study of Tanning Phenotype in a Population of European Ancestry.
Nan H, Kraft P, Qureshi AA, Guo Q, Chen C, Hankinson SE, Hu FB, Thomas G, Hoover RN, Chanock S, Hunter DJ, Han J.
J Invest Dermatol. 2009 Apr 2
Abstract: We conducted a multistage genome-wide association study (GWAS) of tanning response after exposure to sunlight in over 9,000 men and women of European ancestry who live in the United States. An initial analysis of 528,173 single-nucleotide polymorphisms (SNPs) genotyped on 2,287 women identified LOC401937 (rs966321) on chromosome 1 as a novel locus highly associated with tanning ability, and we confirmed this association in 870 women controls from a skin cancer case-control study with joint P-value=1.6 x 10(-9). We further genotyped this SNP in two subsequent replication studies (one with 3,750 women and the other with 2,405 men). This association was not replicated in either of these two studies. We found that several SNPs reaching the genome-wide significance level are located in or adjacent to the loci previously known as pigmentation genes: MATP, IRF4, TYR, OCA2, and MC1R. Overall, these tanning ability-related loci are similar to the hair color-related loci previously reported in the GWAS of hair color.
Blue eyes follow-up
According to this paper, blue eyes in N. European humans is predicted very well by these variants:
HERC2 rs12913832
OCA2 rs1800407
SLC24A4 rs12896399
SLC45A2 rs16891982
TYR rs1393350
IRF4 rs12203592
It might be interesting to look at the five latter ones in the blue-eyed lemur... although, it looks like only one SNP (in HERC2, I assume) is sufficient to explain the majority of the variation in humans.
Thursday, April 02, 2009
Convergent evolution via different genetic mechanism: blue eyes in lemurs and humans
I guess we are to assume that the blue eye color is the same in humans and lemurs. From looking at some pictures, the color appears pretty similar. Haven't read the paper, but are all cases of blue eye color in humans explained by the same variation in HERC2? I might have a post on this somewhere.
I suppose we shouldn't be too surprised by this finding, given what we know about convergence of light skin color evolution in Europeans and E. Asians via different genes.
Brief communication: Blue eyes in lemurs and humans: Same phenotype, different genetic mechanism.
Bradley BJ, Pedersen A, Mundy NI.
Am J Phys Anthropol. 2009 Mar 10.
Almost all mammals have brown or darkly-pigmented eyes (irises), but among primates, there are some prominent blue-eyed exceptions. The blue eyes of some humans and lemurs are a striking example of convergent evolution of a rare phenotype on distant branches of the primate tree. Recent work on humans indicates that blue eye color is associated with, and likely caused by, a single nucleotide polymorphism (rs12913832) in an intron of the gene HERC2, which likely regulates expression of the neighboring pigmentation gene OCA2. This raises the immediate question of whether blue eyes in lemurs might have a similar genetic basis. We addressed this by sequencing the homologous genetic region in the blue-eyed black lemur (Eulemur macaco flavifrons; N = 4) and the closely-related black lemur (Eulemur macaco macaco; N = 4), which has brown eyes. We then compared a 166-bp segment corresponding to and flanking the human eye-color-associated region in these lemurs, as well as other primates (human, chimpanzee, orangutan, macaque, ring-tailed lemur, mouse lemur). Aligned sequences indicated that this region is strongly conserved in both Eulemur macaco subspecies as well as the other primates (except blue-eyed humans). Therefore, it is unlikely that this regulatory segment plays a major role in eye color differences among lemurs as it does in humans. Although convergent phenotypes can sometimes come about via the same or similar genetic changes occurring independently, this does not seem to be the case here, as we have shown that the genetic basis of blue eyes in lemurs differs from that of humans.
Saturday, March 07, 2009
Disentangling genetic signatures of natural selection and demography among Native Americans and W. Beringians
Haplotypic background of a private allele at high frequency in the Americas.
Schroeder KB, Jakobsson M, Crawford MH, Schurr TG, Boca SM, Conrad DF, Tito RY, Osipova LP, Tarskaia LA, Zhadanov SI, Wall JD, Pritchard JK, Malhi RS, Smith DG, Rosenberg NA.
Mol Biol Evol. 2009 Feb 17. [Epub ahead of print]
Schroeder KB, Jakobsson M, Crawford MH, Schurr TG, Boca SM, Conrad DF, Tito RY, Osipova LP, Tarskaia LA, Zhadanov SI, Wall JD, Pritchard JK, Malhi RS, Smith DG, Rosenberg NA.
Mol Biol Evol. 2009 Feb 17. [Epub ahead of print]
Abstract: Recently, the observation of a high-frequency private allele, the 9-repeat allele at microsatellite D9S1120, in all sampled Native American and Western Beringian populations has been interpreted as evidence that all modern Native Americans descend primarily from a single founding population. However, this inference assumed that all copies of the 9-repeat allele were identical by descent and that the geographic distribution of this allele had not been influenced by natural selection. To investigate whether these assumptions are satisfied, we genotyped 34 SNPs across approximately 500 kilobases (kb) around D9S1120 in 21 Native American and Western Beringian populations and 54 other worldwide populations. All chromosomes with the 9-repeat allele share the same haplotypic background in the vicinity of D9S1120, suggesting that all sampled copies of the 9-repeat allele are identical by descent. Ninety-one percent of these chromosomes share the same 76.26 kb haplotype, which we call the "American Modal Haplotype" (AMH). Three observations lead us to conclude that the high frequency and widespread distribution of the 9-repeat allele are unlikely to be the result of positive selection: 1) aside from its association with the 9-repeat allele, the AMH does not have a high frequency in the Americas, 2) the AMH is not unusually long for its frequency compared to other haplotypes in the Americas, and 3) in Latin American mestizo populations, the proportion of Native American ancestry at D9S1120 is not unusual compared to that observed at other genomewide microsatellites. Using a new method for estimating the time to the most recent common ancestor (MRCA) of all sampled copies of an allele on the basis of an estimate of the length of the genealogy descended from the MRCA, we calculate the mean time to the MRCA of the 9-repeat allele to be between 7,325 and 39,900 years, depending on the demographic model used. The results support the hypothesis that all modern Native Americans and Western Beringians trace a large portion of their ancestry to a single founding population which may have been isolated from other Asian populations prior to expanding into the Americas.
Discovery of rare variants in regions identified by GWASs
P-ter at GNXP and Dan at Genetic Future discuss what seems to be an interesting and important recently published paper in Science (abstract below) that finds four rare variants with larger-than-usual effects for Type-1 Diabetes. The last line of the abstract pretty much says it all. The evolutionary interpretation is of course very interesting, especially since the rare variants reduce risk. Dan talks about this.
Sergey Nejentsev, Neil Walker , David Riches, Michael Egholm, John A. Todd
Science DOI: 10.1126/science.1167728
Rare Variants of IFIH1, a Gene Implicated in Antiviral Responses, Protect Against Type 1 Diabetes
Sergey Nejentsev, Neil Walker , David Riches, Michael Egholm, John A. Todd
Science DOI: 10.1126/science.1167728
Abstract: Genome-wide association studies (GWAS) are widely used to map genomic regions contributing to common human diseases, but they often do not identify the precise causative genes and sequence variants. To identify causative type 1 diabetes (T1D) variants, we resequenced exons and splice sites of ten candidate genes in pools of DNA from 480 patients and 480 controls and tested their disease association in over 30,000 subjects. We discovered four rare variants that lowered T1D risk independently of each other (OR = 0.51 – 0.74; P = 1.3 x 10⁻³ – 2.1 x 10⁻¹⁶) in IFIH1, a gene located in a region previously associated with T1D by GWAS. These variants are predicted to alter the expression and structure of IFIH1 (MDA5), a cytoplasmic helicase that mediates induction of interferon response to viral RNA. This firmly establishes the role of IFIH1 in T1D and demonstrates that resequencing studies can pinpoint disease-causing genes in genomic regions initially identified by GWAS.
Friday, March 06, 2009
Introgression of black coat color into wolves from dogs
Molecular and Evolutionary History of Melanism in North American Gray Wolves
Tovi M. Anderson, Bridgett M. vonHoldt, Sophie I. Candille, Marco Musiani, Claudia Greco, Daniel R. Stahler, Douglas W. Smith, Badri Padhukasahasram, Ettore Randi, Jennifer A. Leonard, Carlos D. Bustamante, Elaine A. Ostrander, Hua Tang, Robert K. Wayne, Gregory S. Barsh
Science 6 March 2009: Vol. 323. no. 5919, pp. 1339 - 1343
Abstract: Morphological diversity within closely related species is an essential aspect of evolution and adaptation. Mutations in the Melanocortin 1 receptor (Mc1r) gene contribute to pigmentary diversity in natural populations of fish, birds, and many mammals. However, melanism in the gray wolf, Canis lupus, is caused by a different melanocortin pathway component, the K locus, that encodes a beta-defensin protein that acts as an alternative ligand for Mc1r. We show that the melanistic K locus mutation in North American wolves derives from past hybridization with domestic dogs, has risen to high frequency in forested habitats, and exhibits a molecular signature of positive selection. The same mutation also causes melanism in the coyote, Canis latrans, and in Italian gray wolves, and hence our results demonstrate how traits selected in domesticated species can influence the morphological diversity of their wild relatives.
Wednesday, February 25, 2009
Predictive ability of SNPs for prostate cancer
Utility of Incorporating Genetic Variants for the Early Detection of Prostate Cancer
Robert K. Nam, William W. Zhang, John Trachtenberg, Arun Seth, Laurence H. Klotz, Aleksandra Stanimirovic, Sanoj Punnen, Vasundara Venkateswaran, Ants Toi, D. Andrew Loblaw, Linda Sugar, Katherine A. Siminovitch, Steven A. Narod
Clinical Cancer Research, 10.1158/1078-0432
Abstract Purpose: Several single nucleotide polymorphisms (SNP) have been associated with the risk of prostate cancer. The clinical utility of using SNPs in the early detection of prostate cancer has not been evaluated.Experimental Design: We examined a panel of 25 SNPs from candidate genes and chromosomal regions in 3,004 unselected men who were screened for prostate cancer using serum prostate-specific antigen (PSA) and digital rectal examination. All underwent a prostate biopsy. We evaluated the ability of these SNPs to help predict the presence of prostate cancer at biopsy.Results: Of the 3,004 patients, 1,389 (46.2%) were found to have prostate cancer. Fifteen of the 25 SNPs studied were significantly associated with prostate cancer (P = 0.02-7 x 10⁻⁸). We selected a combination of 4 SNPs with the best predictive value for further study. After adjusting for other predictive factors, the odds ratio for patients with all four of the variant genotypes compared with men with no variant genotype was 5.1 (95% confidence interval, 1.6-16.5; P = 0.006). When incorporated into a nomogram, genotype status contributed more significantly than PSA, family history, ethnicity, urinary symptoms, and digital rectal examination (area under the curve = 0.74). The positive predictive value of the PSA test ranged from 42% to 94% depending on the number of variant genotypes carried (P = 1 x 10-15).Conclusions: SNP genotyping can be used in a clinical setting for the early detection of prostate cancer in a nomogram approach and by improving the positive predictive value of the PSA test.
Wednesday, February 18, 2009
Teaching material for evolution by natural selection
In honor of Darwin's recent 200th birthday, Nature came up with a list of 15 Evolutionary Gems. My major complaint, and a pretty major one at that, is that none of the examples are related to human evolution - something that students could probably relate to a bit more easily, and might increase the chance that they actually pay attention.
Basically, what I'm getting at: Why is lactase persistence not there? or the several other well-documented and clear cut examples from human evolution?
Basically, what I'm getting at: Why is lactase persistence not there? or the several other well-documented and clear cut examples from human evolution?
Elucidating the genetics of eye color
Getting closer to predicting not-so-complex "complex" phenotypes from genetic data?
Eye and skin color are great model phenotypes with which we can learn about the genetics of complex traits, as I've often argued. Here's another step in the process.
Interactions Between HERC2, OCA2 and MC1R May Influence Human Pigmentation Phenotype.
Branicki W, Brudnik U, Wojas-Pelc A.
Ann Hum Genet. 2009 Feb 4. [Epub ahead of print]
Abstract: Human pigmentation is a polygenic trait which may be shaped by different kinds of gene-gene interactions. Recent studies have revealed that interactive effects between HERC2 and OCA2 may be responsible for blue eye colour determination in humans. Here we performed a population association study, examining important polymorphisms within the HERC2 and OCA2 genes. Furthermore, pooling these results with genotyping data for MC1R, ASIP and SLC45A2 obtained for the same population sample we also analysed potential genetic interactions affecting variation in eye, hair and skin colour. Our results confirmed the association of HERC2 rs12913832 with eye colour and showed that this SNP is also significantly associated with skin and hair colouration. It is also concluded that OCA2 rs1800407 is independently associated with eye colour. Finally, using various approaches we were able to show that there is an interaction between MC1R and HERC2 in determination of skin and hair colour in the studied population sample.
Saturday, January 24, 2009
Genetics of lipid profiles in European and African Americans
Genetic Differences between the Determinants of Lipid Profile Phenotypes in African and European Americans: The Jackson Heart Study
Rahul C. Deo, David Reich, Arti Tandon, Ermeg Akylbekova, Nick Patterson, Alicja Waliszewska, Sekar Kathiresan, Daniel Sarpong, Herman A. Taylor, Jr., James G. Wilson
PLoS Genetics 5(1): e1000342.
Rahul C. Deo, David Reich, Arti Tandon, Ermeg Akylbekova, Nick Patterson, Alicja Waliszewska, Sekar Kathiresan, Daniel Sarpong, Herman A. Taylor, Jr., James G. Wilson
PLoS Genetics 5(1): e1000342.
Abstract: Genome-wide association analysis in populations of European descent has recently found more than a hundred genetic variants affecting risk for common disease. An open question, however, is how relevant the variants discovered in Europeans are to other populations. To address this problem for cardiovascular phenotypes, we studied a cohort of 4,464 African Americans from the Jackson Heart Study (JHS), in whom we genotyped both a panel of 12 recently discovered genetic variants known to predict lipid profile levels in Europeans and a panel of up to 1,447 ancestry informative markers allowing us to determine the African ancestry proportion of each individual at each position in the genome. Focusing on lipid profiles—HDL-cholesterol (HDL-C), LDL-cholesterol (LDL-C), and triglycerides (TG)—we identified the lipoprotein lipase (LPL) locus as harboring variants that account for interethnic variation in HDL-C and TG. In particular, we identified a novel common variant within LPL that is strongly associated with TG (p = 2.7×10⁻⁶) and explains nearly 1% of the variability in this phenotype, the most of any variant in African Americans to date. Strikingly, the extensively studied “gain-of-function” S447X mutation at LPL, which has been hypothesized to be the major determinant of the LPL-TG genetic association and is in trials for human gene therapy, has a significantly diminished strength of biological effect when it is found on a background of African rather than European ancestry. These results suggest that there are other, yet undiscovered variants at the locus that are truly causal (and are in linkage disequilibrium with S447X) or that work synergistically with S447X to modulate TG levels. Finally, we find systematically lower effect sizes for the 12 risk variants discovered in European populations on the African local ancestry background in JHS, highlighting the need for caution in the use of genetic variants for risk assessment across different populations.
Friday, January 09, 2009
Gene importance and evolutionary rate
Why Is the Correlation between Gene Importance and Gene Evolutionary Rate So Weak?
Zhi Wang, Jianzhi Zhang
PLoS Genetics 5(1): e1000329.
Zhi Wang, Jianzhi Zhang
PLoS Genetics 5(1): e1000329.
Abstract: One of the few commonly believed principles of molecular evolution is that functionally more important genes (or DNA sequences) evolve more slowly than less important ones. This principle is widely used by molecular biologists in daily practice. However, recent genomic analysis of a diverse array of organisms found only weak, negative correlations between the evolutionary rate of a gene and its functional importance, typically measured under a single benign lab condition. A frequently suggested cause of the above finding is that gene importance determined in the lab differs from that in an organism's natural environment. Here, we test this hypothesis in yeast using gene importance values experimentally determined in 418 lab conditions or computationally predicted for 10,000 nutritional conditions. In no single condition or combination of conditions did we find a much stronger negative correlation, which is explainable by our subsequent finding that always-essential (enzyme) genes do not evolve significantly more slowly than sometimes-essential or always-nonessential ones. Furthermore, we verified that functional density, approximated by the fraction of amino acid sites within protein domains, is uncorrelated with gene importance. Thus, neither the lab-nature mismatch nor a potentially biased among-gene distribution of functional density explains the observed weakness of the correlation between gene importance and evolutionary rate. We conclude that the weakness is factual, rather than artifactual. In addition to being weakened by population genetic reasons, the correlation is likely to have been further weakened by the presence of multiple nontrivial rate determinants that are independent from gene importance. These findings notwithstanding, we show that the principle of slower evolution of more important genes does have some predictive power when genes with vastly different evolutionary rates are compared, explaining why the principle can be practically useful despite the weakness of the correlation.
Friday, January 02, 2009
Genetic information doesn't add much for predicting diabetes
Predicting type 2 diabetes based on polymorphisms from genome-wide association studies: a population-based study.
van Hoek M, Dehghan A, Witteman JC, van Duijn CM, Uitterlinden AG, Oostra BA, Hofman A, Sijbrands EJ, Janssens AC.
Diabetes. 2008 Nov;57(11):3122-8.
van Hoek M, Dehghan A, Witteman JC, van Duijn CM, Uitterlinden AG, Oostra BA, Hofman A, Sijbrands EJ, Janssens AC.
Diabetes. 2008 Nov;57(11):3122-8.
OBJECTIVE: Prediction of type 2 diabetes based on genetic testing might improve identification of high-risk subjects. Genome-wide association (GWA) studies identified multiple new genetic variants that associate with type 2 diabetes. The predictive value of genetic testing for prediction of type 2 diabetes in the general population is unclear. RESEARCH DESIGN AND METHODS: We investigated 18 polymorphisms from recent GWA studies on type 2 diabetes in the Rotterdam Study, a prospective, population-based study among homogeneous Caucasian individuals of 55 years and older (genotyped subjects, n = 6,544; prevalent cases, n = 686; incident cases during follow-up, n = 601; mean follow-up 10.6 years). The predictive value of these polymorphisms was examined alone and in addition to clinical characteristics using logistic and Cox regression analyses. The discriminative accuracy of the prediction models was assessed by the area under the receiver operating characteristic curves (AUCs). RESULTS: Of the 18 polymorphisms, the ADAMTS9, CDKAL1, CDKN2A/B-rs1412829, FTO, IGF2BP2, JAZF1, SLC30A8, TCF7L2, and WFS1 variants were associated with type 2 diabetes risk in our population. The AUC was 0.60 (95% CI 0.57-0.63) for prediction based on the genetic polymorphisms; 0.66 (0.63-0.68) for age, sex, and BMI; and 0.68 (0.66-0.71) for the genetic polymorphisms and clinical characteristics combined. CONCLUSIONS: We showed that 9 of 18 well-established genetic risk variants were associated with type 2 diabetes in a population-based study. Combining genetic variants has low predictive value for future type 2 diabetes at a population-based level. The genetic polymorphisms only marginally improved the prediction of type 2 diabetes beyond clinical characteristics.
Tuesday, December 23, 2008
The genetics of skin pigmentation in beach mice from different beaches
A similar story to humans in East Asia and Europe...
The Genetic Basis of Phenotypic Convergence in Beach Mice: Similar Pigment Patterns but Different Genes Cynthia C. Steiner, Holger Römpler, Linda M. Boettger, Torsten Schöneberg and Hopi E. Hoekstra
Molecular Biology and Evolution 2009 26(1):35-45
Abstract: Convergent evolution is a widespread phenomenon seen in diverse organisms inhabiting similar selective environments. However, it is unclear if similar phenotypes are produced by the same or different genes and mutations. Here we analyze the molecular mechanisms underlying convergent pigment pattern among subspecies of the beach mouse (Peromyscus polionotus) inhabiting the Gulf and Atlantic coasts of Florida. In these two geographic regions, separated by more than 300 km, "beach mice" have lighter colored coats than do their mainland counterparts, produced by natural selection for camouflage against the pale coastal sand dunes. We measured color pattern in eight beach mouse subspecies and showed that three of the Gulf Coast subspecies are more phenotypically similar to an Atlantic coast subspecies than to their Gulf Coast neighbors. However, light-colored beach mice do not form a monophyletic group. Previous results implicated a single derived amino acid change in the melanocortin-1 receptor (Mc1r) as a major contributor to pigment pattern in the Gulf Coast beach mice; despite phenotypic similarities, the derived Mc1r allele was not found in the Atlantic coast beach mouse populations. Here we show that Atlantic coast beach mice have high levels of Mc1r polymorphism but they lack unique alleles. Functional assays revealed that single amino acid mutations segregating in Atlantic coast beach mice do not cause any change in Mc1r activity compared with the activity of Mc1r from dark-colored mice. These joint results show that convergent pigment patterns in recently diverged beach mouse subspecies—whose developmental constraints are presumably similar—have evolved through a diversity of genetic mechanisms.
Thursday, December 11, 2008
Tradeoff between pathogen resistance and auto-immune response?
I first read about this in one of the earlier papers on European genetic structure where they mention: "a north/south gradient in the incidence of autoimmune diseases has been noted for several continents," and give a reference to this paper which, if I remember correctly, shows differences in incidence of various diseases across Europe and discusses the potential tradeoff between pathogen resistance and auto-immune disorders.
Dan at Genetic Future points out a recent study where they find that the same genetic factors contribute to both celiac disease and type I diabetes (both auto-immune disorders). The study also finds that those with the delta-32 variant of CCR5 (protective of HIV infection) have a lower risk of type I diabetes and celiac disease.
These findings beg the questions: Given the distribution of these diseases across Europe, shouldn't they have controlled for ancestry stratification?, and more generally: Is there a unifying phenotype that can be measured which maps to the underlying factor contributing to all these auto-immune diseases? Why is this phenotype seemingly more prevalent among Northern Europeans? Is there some kind of tradeoff between between pathogen resistance and auto-immune "disorders"?
I must say I'm still unclear about the evidence, if any, for this tradeoff.
Wednesday, December 10, 2008
Slavery, fetal programming, and low birth weight among African Americans
Low birth weight of contemporary African Americans: An intergenerational effect of slavery?
Grazyna Jasienska
American Journal of Human Biology 2008, 21: 16 - 24
Abstract: The average birth weight in the contemporary African-American population is about 250 g lower than the average birth weight of European Americans. Differences in genetic and socioeconomic factors present between these two groups can explain only part of birth weight variation. I propose a hypothesis that the low birth weight of contemporary African Americans not only results from the difference in present exposure to lifestyle factors known to affect fetal development but also from conditions experienced during the period of slavery. Slaves had poor nutritional status during all stages of life because of the inadequate dietary intake accompanied by high energetic costs of physical work and infectious diseases. The concept of fetal programming suggests that physiology and metabolism including growth and fat accumulation of the developing fetus, and, thus its birth weight, depend on intergenerational signal of environmental quality passed through generations of matrilinear ancestors. I suggest that several generations that have passed since the abolition of slavery in the United States (1865) has not been enough to obliterate the impact of slavery on the current biological and health condition of the African-American population.
Subscribe to:
Posts (Atom)